MedIntegrity IntegtestV8.0 Manufacturer & Supplier
Integtest V8.0 Filter Integrity Tester
Overview
Designed and manufactured by MedIntegrity LLC — the filter integrity tester is built in our own factory, part of a 28-patent product line in pharmaceutical testing equipment.
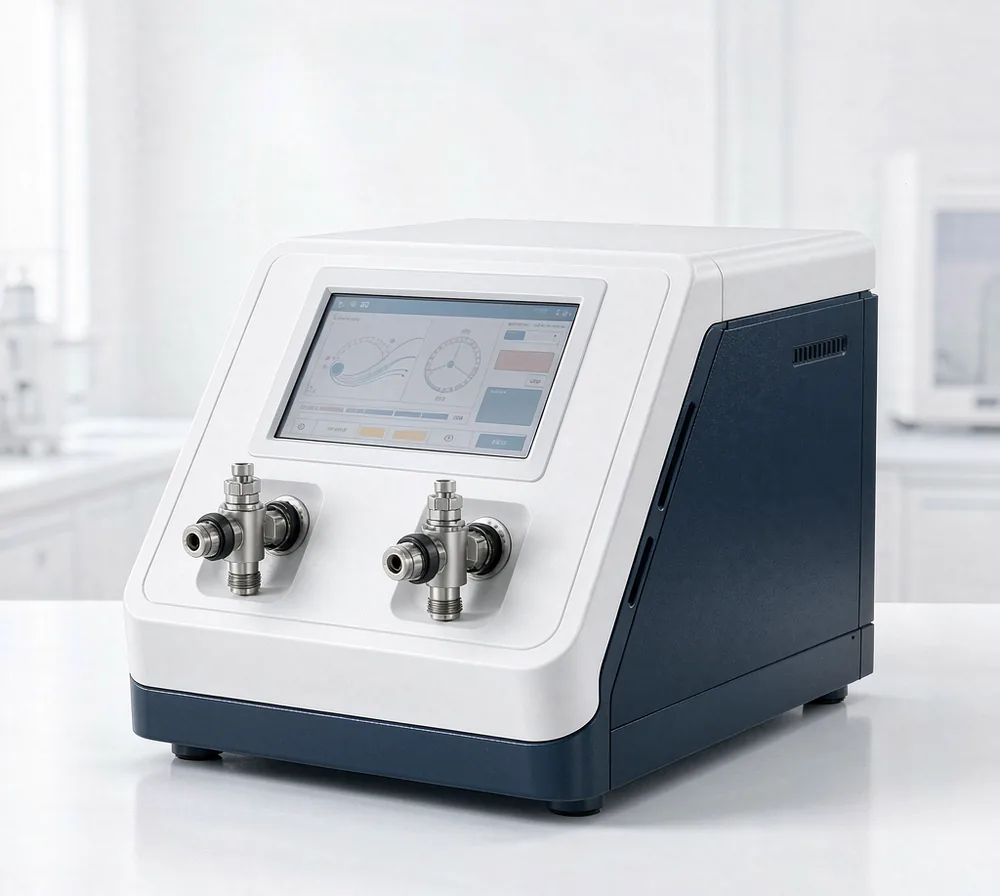
MedIntegrity Filter Integrity Testing Technology Integtest V8.0 Establishing a Contamination Control Strategy The MedIntegrity Integrity measurement Integtest V8.0 systems represent the latest generation of best-in-class integrity test equipment. The instruments have a very sensitive measurement system and are capable of detecting almost all conditions developed in accordance with GAMP guidelines, and offering full compliance with 21 CFR Part 11 is supported by an extensive qualification documentation package, including guides for Installation and Operational Qualification Performance Qualification(IQ/OQ/PQ)MedIntegrity - USANote:Please so kindly understand that the information and specification in this catalogue are subject to change for improvement without notice.If necessary,or any technical question, please contact your MedIntegrity representative What are the Regulations/Guidance for Sterilizing Filter Testing ·Sterilizing Filter"A filter that, when appropriately validated, will remove all microorganisms from a fluid stream, producing a sterile effluent"
Technical Specifications
| Audit trail record storage | ≥5 years storage |
|---|---|
| Number of users | 1000setsOperating System is Linux system |
| Dust and splash level | IP54,Front is IP65Print Function is no data |
| Serial Connection | (4-20)MA,RS485,12V Alarm output (Optional)Language is Chinese/English |
| Weight | 10KG |
Specifications subject to change for improvement without notice. For the latest data sheet or technical questions, contact your MedIntegrity representative.
In practice, the IntegtestV8.0 walks through the method set — bubble point to vacuum decay — and stores the results as records your QC file can use directly. If the filter is sterilizing grade, we'll help set the acceptance limit before you run it.
Technical Documentation
Notes From Our Technical Documentation
Regulations & Guidance for Sterilizing Filter Testing
What are the Regulations/Guidance for Sterilizing Filter Testing 2.1 FDA Guidance for Industry-Sterile Drug Products Produced by Aseptic Processing-Current Good Manufacturing Practice September,2004 ·Sterilizing Filter"A filter that, when appropriately validated, will remove all microorganisms from a fluid stream, producing a sterile effluent" 2.2 EU GMP Annex 1 Sterile Medicinal Products Annex 1 has been revised on numerous occasions (1997, 2008, 2022) with each revision accommodating advances in technology,understanding and industry trendsThe most recent update due for implementation in 2023 contains numerous changes. Many of these aim to address points of ambiguity and adapt language to align with technical and regulatory precedents to achieve greater clarity. The requirement for pre-use, post-sterilization filter integrity testing (PUPSIT) is among the areas that receive adjusted wording. [Heading 4] 2.2.1 Pre-use, post-sterilization integrity testing (EU GMP Annex 1, Sterilizing Grade Filters Used for Aseptic Manufacturing) [Heading 5] 2008: “The integrity of the sterilised filter should be verified before use and should be confirmed immediately after use by an appropriate method such as a bubble point, diffusive flow or ressure hold test. The time taken to filter a known volume of bulk solution and the pressure difference to be used across the filter should be determined during validation and any significant differences from this during routine manufacturing should be noted and investigated.Results of these checks should be included in the batch record. The integrity of critical gas and air vent filters should be confirmed after use. The integrity of other filters should be confirmed at appropriate intervals.”
Technical Guide
Why the measurement side matters
Sterilizing-grade integrity testing lives or dies on small signals. Diffusive flow is a small number measured against instrument drift, and a pressure hold that is slowly drifting tells you things a single point reading never will. The V8.0's measurement system is built for that regime — sensitive enough that a change in the state of the filter is visible in the data before it becomes a problem in the batch record.
The method set is the standard one for sterilizing-grade filtration: bubble point for the largest pore, diffusive flow for the small-pore population and contamination, pressure hold for seal and membrane stability, and vacuum decay for dry filters and gas-side filters. Your procedure picks the method that matches the question, or runs the combination it defines.
Built for the line, not just the lab
The front panel carries an IP65 rating and the instrument body IP54 — and the difference is deliberate. The front is the side that faces the washdown area of the room: dust-tight where it gets splashed, and sealed enough against dust and water jets that routine cleaning is a squeegee and a wipe rather than a shutdown event.
The V8.0 also talks to the plant. The interface set includes 4–20 mA, RS485 and an optional 12 V alarm output — the standard connectors for linking an instrument result into line control or an alarm system. A failed integrity check can trigger an action at the line instead of sitting in a database waiting for someone to notice.
Users, records and the audit trail
The system supports up to 1,000 user accounts on a Linux operating system, with role-based access — the structure 21 CFR Part 11 expects when different people operate, review and release. The operating language is Chinese or English, selected at the instrument.
The audit trail is sized for the long view: storage of five years or more. That matches the retention expectations of most GMP quality systems, and retrieval is by time, so pulling everything for a campaign window is a search, not a reconstruction.
Delivery includes the qualification documentation package — guides for IQ, OQ and PQ — developed in accordance with GAMP guidelines with full 21 CFR Part 11 compliance support. The instrument and the documentation arrive together, because in an audit they get read together.
The PUPSIT requirement, met with one instrument
EU GMP Annex 1 has been revised in 1997, 2008 and 2022, and the 2023 implementation tightened the wording around one point that matters most in routine aseptic filtration: pre-use, post-sterilization integrity testing. The integrity of the sterilizing filter is verified before use and confirmed immediately after use — by a bubble point, diffusive flow or pressure hold test — with the results carried into the batch record, and the integrity of critical gas and air vent filters confirmed after use as well.
The V8.0's method set is built for exactly that pattern: the same instrument runs the check before the filter goes to service and the confirmation after it has done its work, and because every run is on the same method and the same measurement baseline, the before and after numbers are directly comparable. When routine results drift from the values determined during validation, the data shows it — which is what the Annex expects you to note and investigate.
Reading small signals on the line
The practical difficulty of sterilizing-grade integrity testing is that the important results are small. A diffusive flow number is a low, steady value measured against the instrument's own drift, and a pressure hold that creeps downward over the test window is a slower signal than a single point reading would ever show. An instrument that only reports a headline number can miss the trend, and the trend is usually where the filter is telling you something. The V8.0's measurement side is built to see that: the resolution and stability are sized so that a change in the state of the membrane shows up in the data before it becomes a line stop.
That is why the method set matters as a set. Bubble point tells you the largest pore is where it should be; diffusive flow tells you the small-pore population and the contamination loading are holding; pressure hold tells you the seal and the membrane are stable across the cycle; vacuum decay covers the dry and gas-side filters in the same loop. Run together on the schedule your procedure defines, they turn a single pass/fail into a documented picture of the filter's state across its working life.
Frequently Asked Questions
What do the IP ratings cover?
IP54 for the instrument body and IP65 on the front panel — protection against dust, and against splashing and water jets on the side that faces routine washdown.
How long is the audit trail retention?
Five years or more, per the specification, with up to 1,000 user accounts on a Linux system and role-based access.
Can it connect to plant systems?
Yes — 4–20 mA, RS485 and an optional 12 V alarm output, the standard links for connecting an integrity result into line control or an alarm system.
Which methods are available?
Bubble point, diffusive flow, pressure hold and vacuum decay, for sterilizing-grade liquid and gas filters per USP <43> and EP 11.0.
What's in the delivery?
The instrument plus the qualification documentation package (IQ/OQ/PQ), developed in accordance with GAMP guidelines with 21 CFR Part 11 support.
Which test methods are available?
Bubble point, diffusive flow, pressure hold and vacuum decay — covering pre-use, post-use and PUPSIT verification for sterilizing-grade liquid filters.
Which pharmacopeial standards does it align with?
USP <43> and EP 11.0 filtration verification, used for pre- and post-sterilization integrity testing of membrane filters.
Can results be documented for audits?
Yes. Results are captured as records that support audit documentation, with 21 CFR Part 11 support across our instruments.
Need a quote or technical advice?
Tell us your application — our U.S. team responds with product selection, documentation and pricing.
Request a Quote